2026-10-07 スタンフォード大学
<関連情報>
- https://news.stanford.edu/stories/2026/10/gut-microbiome-human-migration
- https://www.nature.com/articles/s41586-026-11106-1
- https://www.cell.com/cell/fulltext/S0092-8674(23)00597-4
- https://www.nature.com/articles/s41467-020-17541-6
人類とともに消滅しつつある腸内微生物が先史時代に世界規模で移動した Prehistoric global migration of vanishing gut microbes with humans
Matthew M. Carter, Zhiru Liu, Matthew R. Olm, Melanie Martin, Daniel D. Sprockett, Parsa Ghadermazi, Benjamin C. Trumble, Hillard Kaplan, Jonathan Stieglitz, Daniel Eid Rodriguez, David A. Relman, Erica D. Sonnenburg, Michael Gurven, Benjamin H. Good & Justin L. Sonnenburg
Nature Published:07 October 2026
DOI:https://doi.org/10.1038/s41586-026-11106-1
Abstract
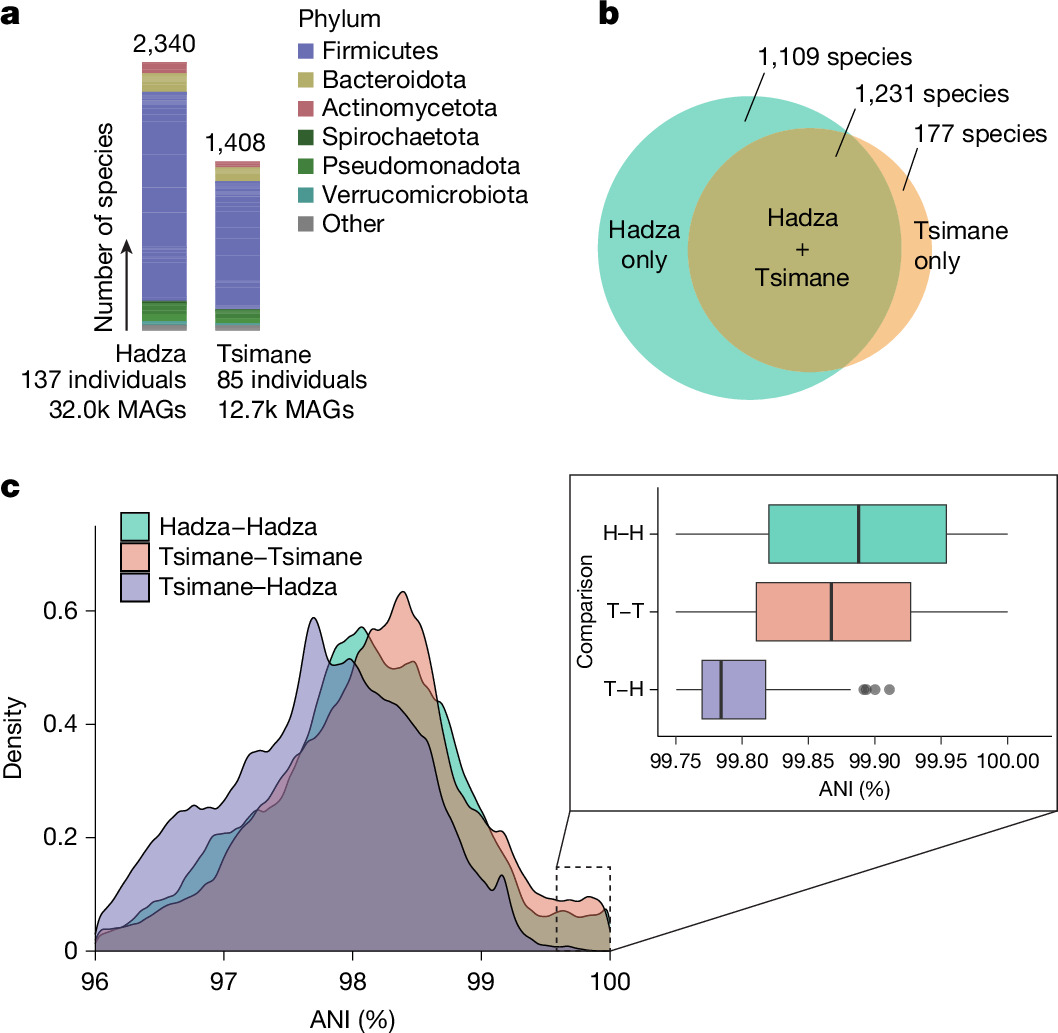
The gut microbiome is crucial for health and is affected strongly by lifestyle1. Many microorganisms commonly found in non-industrialized populations are disappearing or have become extinct in industrialized populations2,3,4,5,6. Studying which microorganisms have been long-term residents of the human gut and may have co-evolved with humans2,7,8 could provide insights into how microbial biodiversity loss affects human health. However, the genetic complexities of microbial evolution and the plasticity of gut microbiome composition have made it challenging to resolve the evolutionary history of these long-term associations. Here we performed deep metagenomic sequencing of the Tsimane horticulturalists of Bolivia and compared their gut microbiomes with those of the Hadza hunter-gatherers of Tanzania3. These two populations, whose ancestors have been separated for tens of thousands of years, share 1,231 microbial species, most of which are rare in or absent from industrialized populations. Population genetic analyses of 636 of the shared species revealed patterns of microbial divergence and gene flow consistent with prehistoric human co-migration, with estimated split times that approximately align with human migration out of Africa and into the Americas. Our findings indicate that a diverse gut microbiome co-migrated with humans worldwide and has persisted over millennia. However, many of these species are now vanishing from industrialized populations and the consequences for human health remain uncertain.
ハザ族の狩猟採集民の超高深度ゲノムシーケンス解析により、消滅しつつある腸内微生物が発見される Ultra-deep sequencing of Hadza hunter-gatherers recovers vanishing gut microbes
Matthew M. Carter ∙ Matthew R. Olm ∙ Bryan D. Merrill ∙ … ∙ Aashish R. Jha ∙ Erica D. Sonnenburg ∙ Justin L. Sonnenburg
Cell Published:June 21, 2023
DOI:https://doi.org/10.1016/j.cell.2023.05.046
Highlights
- The largest set of gut microbiome sequencing data from a hunter-gatherer population
- Assembly of thousands of novel human gut bacteria, archaea, eukaryotes, and phages
- Identification of distinct functions in lifestyle-associated VANISH and BloSSUM taxa
- Extensive non-kin strain sharing associated with a unique Hadza social structure
Summary
The gut microbiome modulates immune and metabolic health. Human microbiome data are biased toward industrialized populations, limiting our understanding of non-industrialized microbiomes. Here, we performed ultra-deep metagenomic sequencing on 351 fecal samples from the Hadza hunter-gatherers of Tanzania and comparative populations in Nepal and California. We recovered 91,662 genomes of bacteria, archaea, bacteriophages, and eukaryotes, 44% of which are absent from existing unified datasets. We identified 124 gut-resident species vanishing in industrialized populations and highlighted distinct aspects of the Hadza gut microbiome related to in situ replication rates, signatures of selection, and strain sharing. Industrialized gut microbes were found to be enriched in genes associated with oxidative stress, possibly a result of microbiome adaptation to inflammatory processes. This unparalleled view of the Hadza gut microbiome provides a valuable resource, expands our understanding of microbes capable of colonizing the human gut, and clarifies the extensive perturbation induced by the industrialized lifestyle.
ボリビア・アマゾンのチマネ族園芸農民における微生物叢の構成、構造、および動態 Microbiota assembly, structure, and dynamics among Tsimane horticulturalists of the Bolivian Amazon
Daniel D. Sprockett, Melanie Martin, Elizabeth K. Costello, Adam R. Burns, Susan P. Holmes, Michael D. Gurven & David A. Relman
Nature Communications Published:29 July 2020
DOI:https://doi.org/10.1038/s41467-020-17541-6
Abstract
Selective and neutral forces shape human microbiota assembly in early life. The Tsimane are an indigenous Bolivian population with infant care-associated behaviors predicted to increase mother-infant microbial dispersal. Here, we characterize microbial community assembly in 47 infant-mother pairs from six Tsimane villages, using 16S rRNA gene amplicon sequencing of longitudinal stool and tongue swab samples. We find that infant consumption of dairy products, vegetables, and chicha (a fermented drink inoculated with oral microbes) is associated with stool microbiota composition. In stool and tongue samples, microbes shared between mothers and infants are more abundant than non-shared microbes. Using a neutral model of community assembly, we find that neutral processes alone explain the prevalence of 79% of infant-colonizing microbes, but explain microbial prevalence less well in adults from river villages with more regular access to markets. Our results underscore the importance of neutral forces during microbiota assembly. Changing lifestyle factors may alter traditional modes of microbiota assembly by decreasing the role of neutral processes.