2026-10-10 中国科学院(CAS)
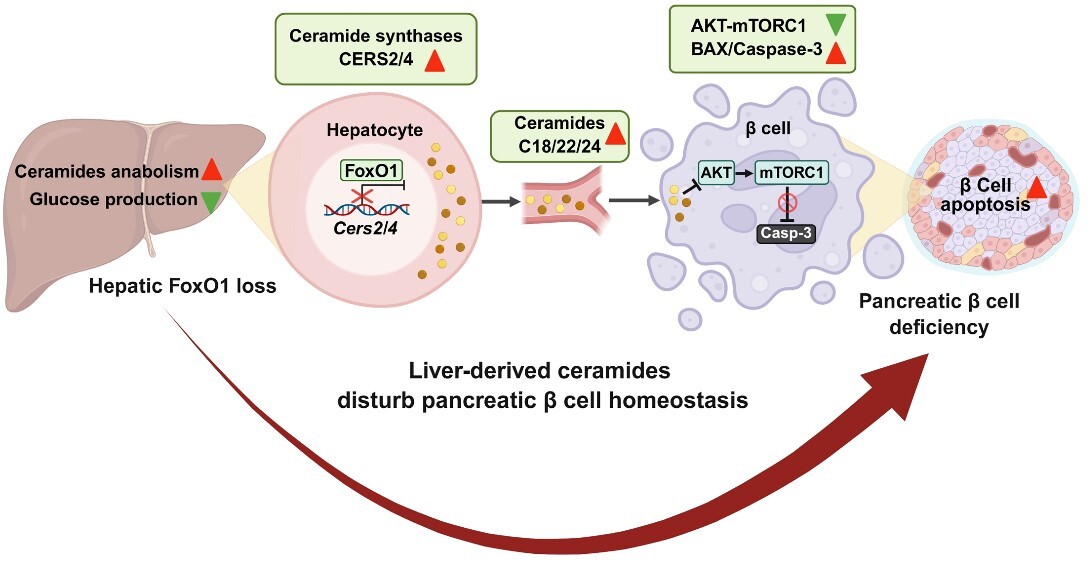
Hepatic FoxO1 loss induces pancreatic β cell deficiency mainly via liver-derived C18/22/24 ceramides. (Image by IHB)
<関連情報>
- https://english.cas.cn/newsroom/research-news/202610/t20261010_1202179.shtml
- https://www.sciencedirect.com/science/article/abs/pii/S2451945626003302
肝臓におけるFoxO1の欠損は、肝臓由来のC18/22/24セラミドを介して膵臓β細胞機能不全を引き起こす Hepatic FoxO1 loss leads to pancreatic β cell deficiency via liver-derived C18/22/24 ceramides
Yulong Gong, Qisheng Lu, Xuehui Li, Guoli Han, Xiaochen Ma, Yulong Liu, Longwei Xi, Chaofan Wang, Gang Zhai, Zhimin Zhang, Haokun Liu, Junyan Jin, Yunxia Yang, Xiaoming Zhu, Zhan Yin, Yonghua Sun, Wenbiao Chen, Shouqi Xie, Dong Han
Cell Chemical Biology Available online: 1 October 2026
DOI:https://doi.org/10.1016/j.chembiol.2026.09.002
Highlights
- Hepatic FoxO1 loss induces pancreatic β cell apoptosis via inter-organ signal
- FoxO1 acts as a transcription repressor for hepatic Cers2/4
- Hepatic FoxO1 loss promotes C18/22/24 ceramide production via elevating Cers2/4
- Liver-derived ceramides contribute to β cell apoptosis by inhibiting AKT-mTORC1
Significance
Organs communicate with each other through secreted signals to coordinate whole-body metabolic homeostasis. In modern lifestyles, hyperglycemia is a key feature of metabolic syndrome, and hepatic FoxO1 has emerged as a promising therapeutic target for lowering blood glucose. However, the potential side effects of inhibiting FoxO1 on other organs remain largely unknown. In this study, we discover that hepatic FoxO1 loss triggers the overproduction of specific lipid messengers—C18/22/24 ceramides—that travel to the pancreatic islets and contribute to β cell apoptosis. These findings reveal a previously unrecognized liver-β cell signaling axis and raise an important concern: therapies designed to inhibit hepatic FoxO1 may inadvertently harm pancreatic β cells, suggesting that combining FoxO1 inhibition with strategies to curb ceramide production could be necessary to preserve β cell integrity while achieving glycemic control.
Summary
Preserving pancreatic β cells is requisite for antihyperglycemic strategy. Hepatic FoxO1 represents a promising therapeutic target. However, the systemic consequences on β cell homeostasis upon hepatic FoxO1 loss and the underlying mechanism remain largely unexplored. Using liver-specific Foxo1 knockdown mouse and transgenic zebrafish models, we uncover that hepatic FoxO1 loss leads to β cell deficiency via triggering apoptosis. Lipidomic and pharmacological analyses identify that hepatic FoxO1 loss increases the liver-derived C18/22/24 ceramides, which contribute to apoptosis by inhibiting the AKT-mTORC1 pathway in β cells. The elevated C18/22/24 ceramide production is due to the upregulated transcription of hepatic ceramide synthase 2/4, while ChIP-seq and luciferase assays further decipher that FoxO1 acts as a transcriptional repressor of Cers2/4. We, therefore, report an inter-organ signal from liver-derived ceramides to pancreatic β cell homeostasis upon hepatic FoxO1 loss. These findings provide insights for therapeutic strategies aimed at preserving β cells and ameliorating hyperglycemia.