2026-10-07 京都大学
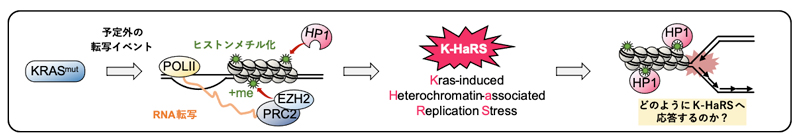
発がん性KRASは転写活動とヘテロクロマチン形成を介してDNA複製を妨害する。細胞がK-HaRSに応答し、適応していく過程は不明だった。
<関連情報>
- https://www.kyoto-u.ac.jp/ja/research-news/2026-10-07-2
- https://www.nature.com/articles/s41467-026-77585-y
ATRによって切り替えられるPrimPolは、発がん性KRAS誘導複製ストレス下でのフォークの減速と再プライミングを調節する PrimPol toggled by ATR modulates fork slowing and repriming under oncogenic KRAS-induced replication stress
Taichi Igarashi, Takaaki Yasuhara & Bunsyo Shiotani
Nature Communications Published:01 October 2026
DOI:https://doi.org/10.1038/s41467-026-77585-y Early provide
Abstract
During the early phases of oncogenic clonal expansion, precancerous cells experience replication stress that threatens both genomic integrity and viability. How replication stress tolerance (RST) mechanisms are dynamically reprogrammed across the different stages of cancer progression remains elusive. Here, we identify PrimPol as a central regulator that executes two fundamentally distinct RST modes in response to oncogenic KRAS-induced heterochromatin-associated replication stress (K-HaRS). Unexpectedly, during acute K-HaRS, an unphosphorylated form of PrimPol cooperates with the fork remodeler SMARCAL1 to actively slow fork progression, stabilizing challenged forks and limiting engagement of error-prone DNA synthesis. As K-HaRS persists, PrimPol undergoes an ATR-dependent functional switch, whereby its phosphorylated form promotes repriming while fork slowing is attenuated, enabling replication to proceed across heterochromatin barriers despite persistent stress. Strikingly, loss of PrimPol enforces chronic reliance on the error-prone Rev1-Polζ-dependent synthesis and compromises cellular fitness under KRAS-driven stress. Thus, these findings establish PrimPol as a context-dependent regulator of RST that, through ATR-dependent toggling of its functional modes, coordinates fork slowing and repriming to safeguard replication fork stability while permitting an optimal compromise of genome stability that would fuel oncogenic clonal evolution.